Nature Chemistry

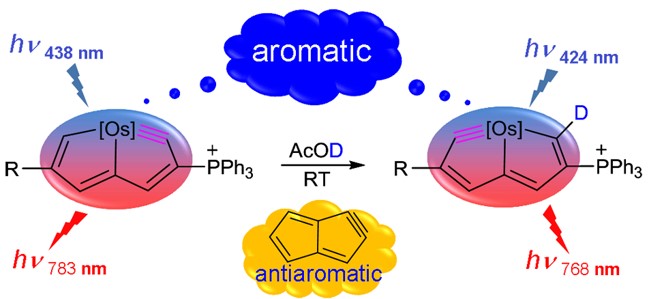
Aromaticity, a cornerstone of chemical stability, is governed by Hückel’s and Baird’s rules for the ground (S0) and triplet excited (T1) states, respectively, yet these canonical rules are challenged by two-state aromaticity (adaptive aromaticity)—a burgeoning frontier in aromaticity research.

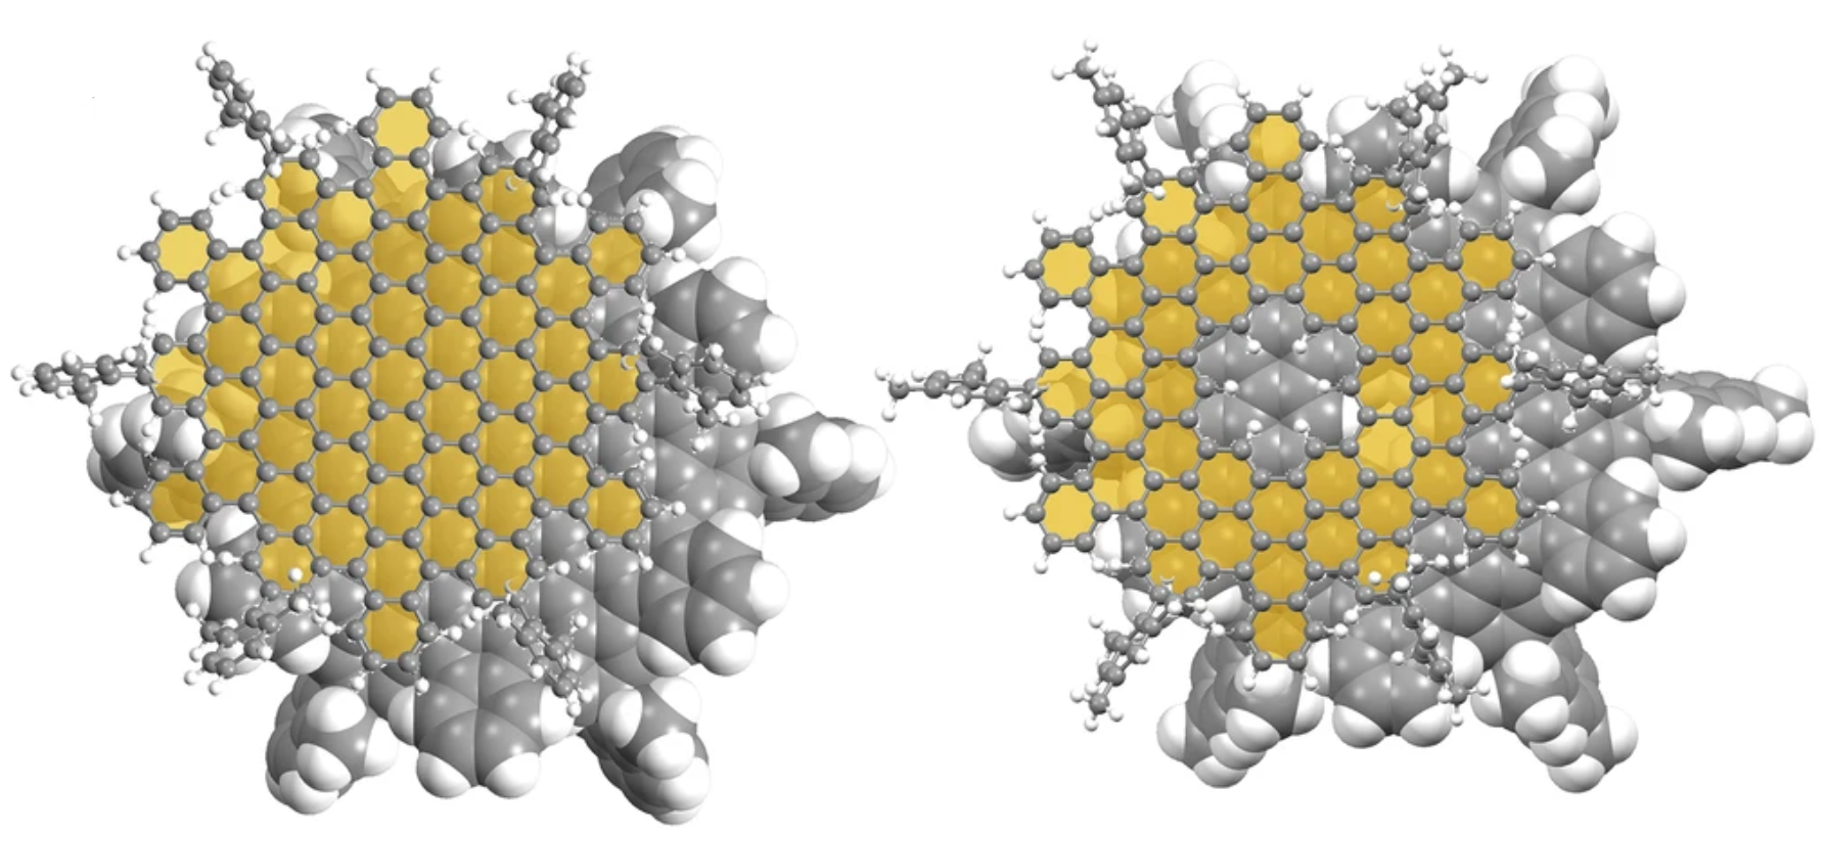
Owing to the lack of precise atomic-level control offered by existing fabrication techniques over the edge structures in bilayer graphene, experimental studies on the edge effects in bilayer graphene structures have so far largely remained unexplored. In this study, two fully cove-edged bilayer nanographenes (CBGs) are synthesized via organic synthesis approach. One of the prepared CBGs contains a cavity defect. The bilayer stacked structures of CBGs are unambiguously characterized.

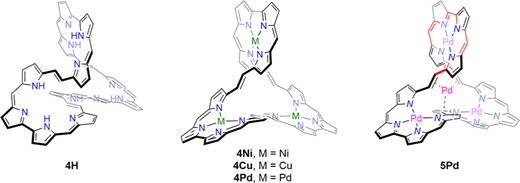
Molecular topology plays a key role in controlling the aromatic or antiaromatic character of cyclic π-conjugated systems. Herein we report synthesis of triply twisted [54]dodecaphyrin(2.1.0.1.2.1.0.1.2.1.0.1) tri-NiII complex 4Ni via reductive cyclization of cyclic tris(dipyrrin) NiII complex 3, which was synthesized by a tri-NiII templation. 4Ni was converted into free base 4H and CuII complex 4Cu.

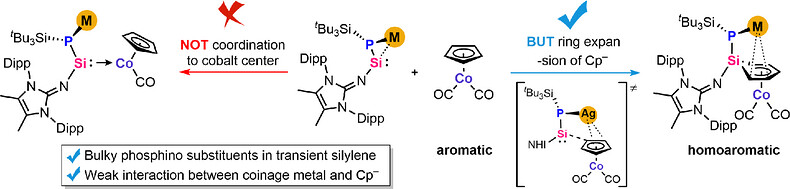
In this study, we report the aromatic C─C bond cleavage of cyclopentadienide by transient imino(metallophosphino)silylenes through the reaction of silylene equivalents with CpCo(CO)2, enabling the transformation of cyclopentadienides into silicon-bridged homocyclopentadienides. The presence of homoaromaticity in these species is substantiated by NICS and ACID analyses.

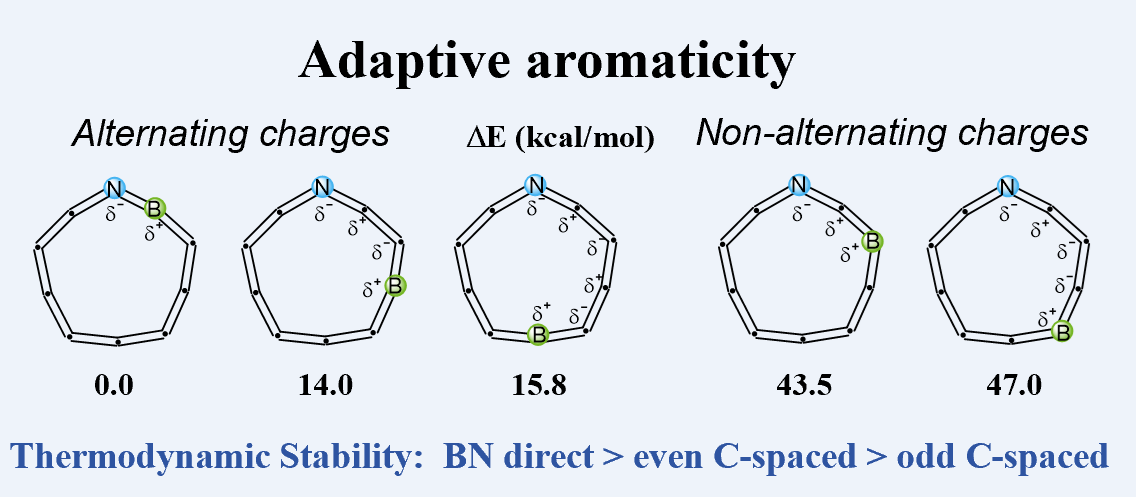
B,N-doped organic analogues can be widely used in photoelectronic materials, medicine and other fields. However, B,N-doped cyclo[n]carbons have not yet been developed. Herein, we report a systematic investigation of the thermodynamic stability and aromaticity of mono B,N-doped cyclo[10]carbon isomers via density functional theory (DFT). Several rules for thermodynamic stability have been proposed. Specifically, rule 1: isomers with B,N atoms separated by an even number of C atoms are more stable than those separated by an odd number of C atoms.

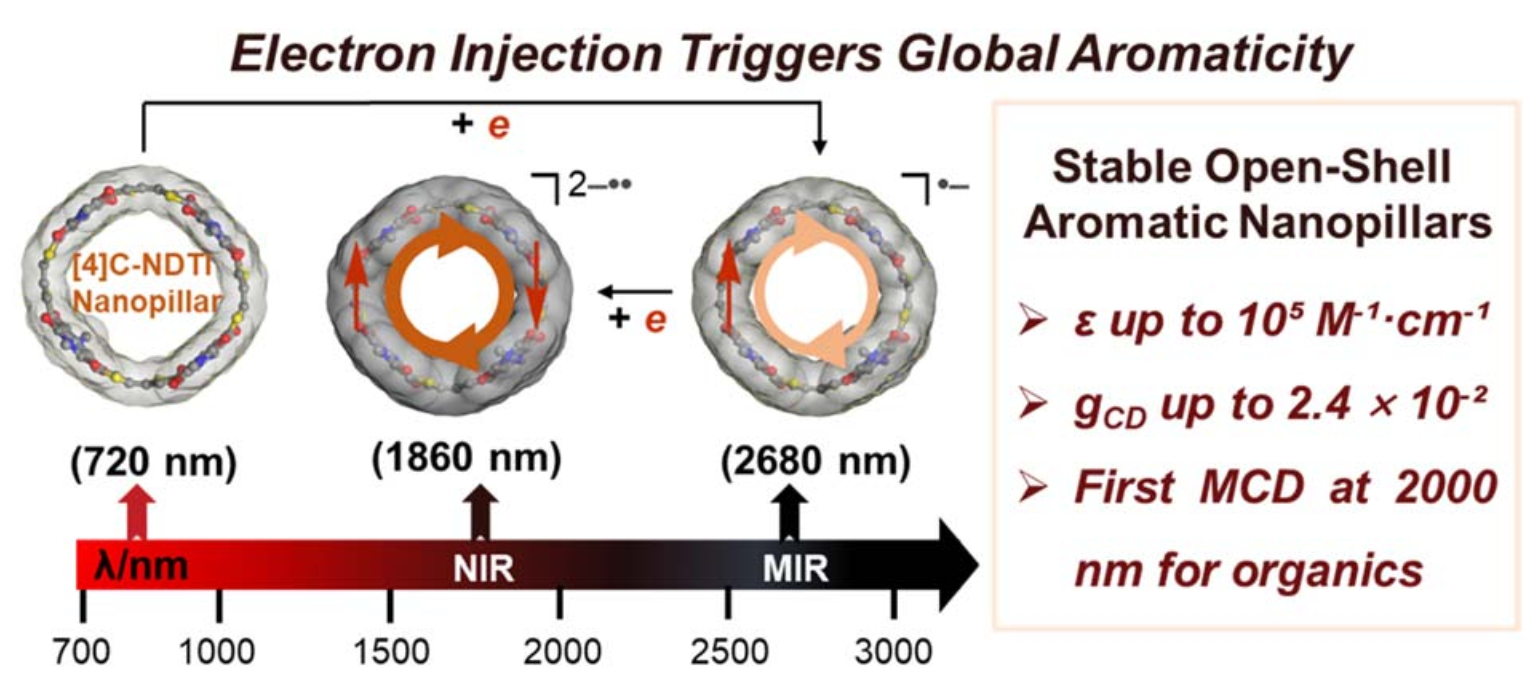
Open-shell species are central to spintronics and infrared optoelectronics, but remain challenging to stabilize in discrete molecular systems. Herein, we report that electron injection into pillar-shaped, radially π-conjugated [4]cyclonaphthodithiophene diimides ([4]C-NDTIs) triggers global aromaticity, yielding radical species with notable stability and optical properties.

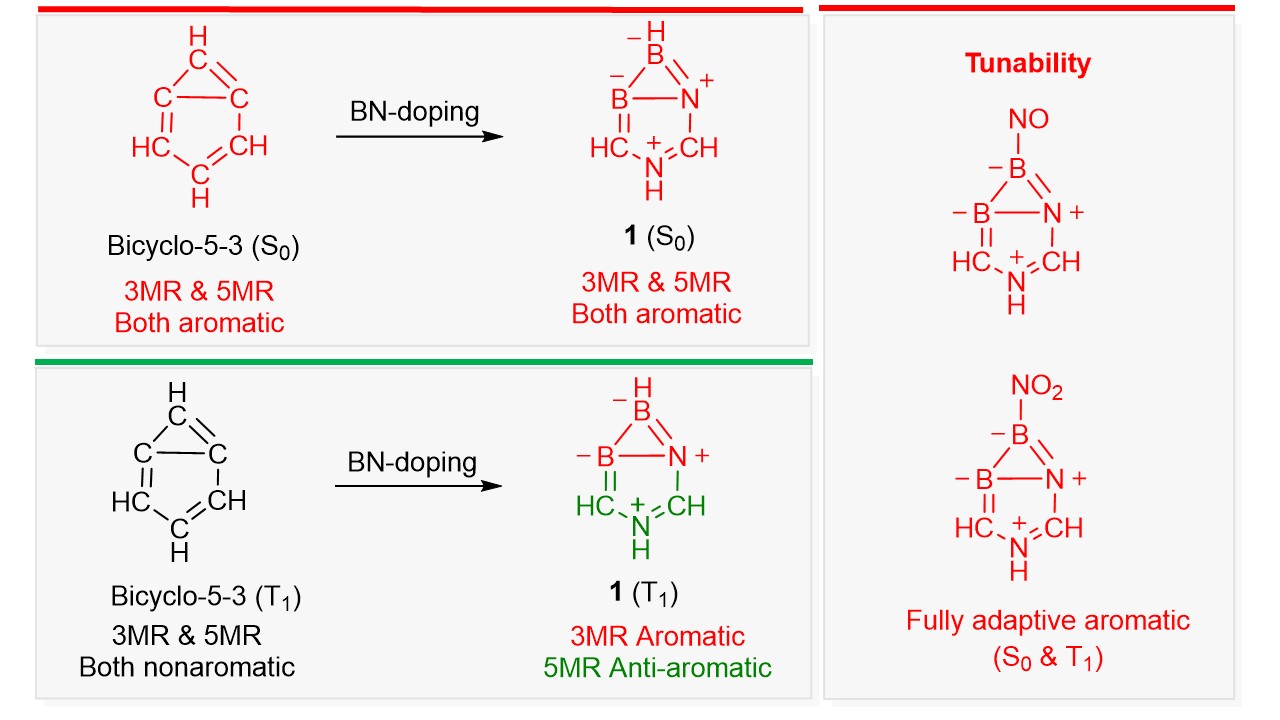
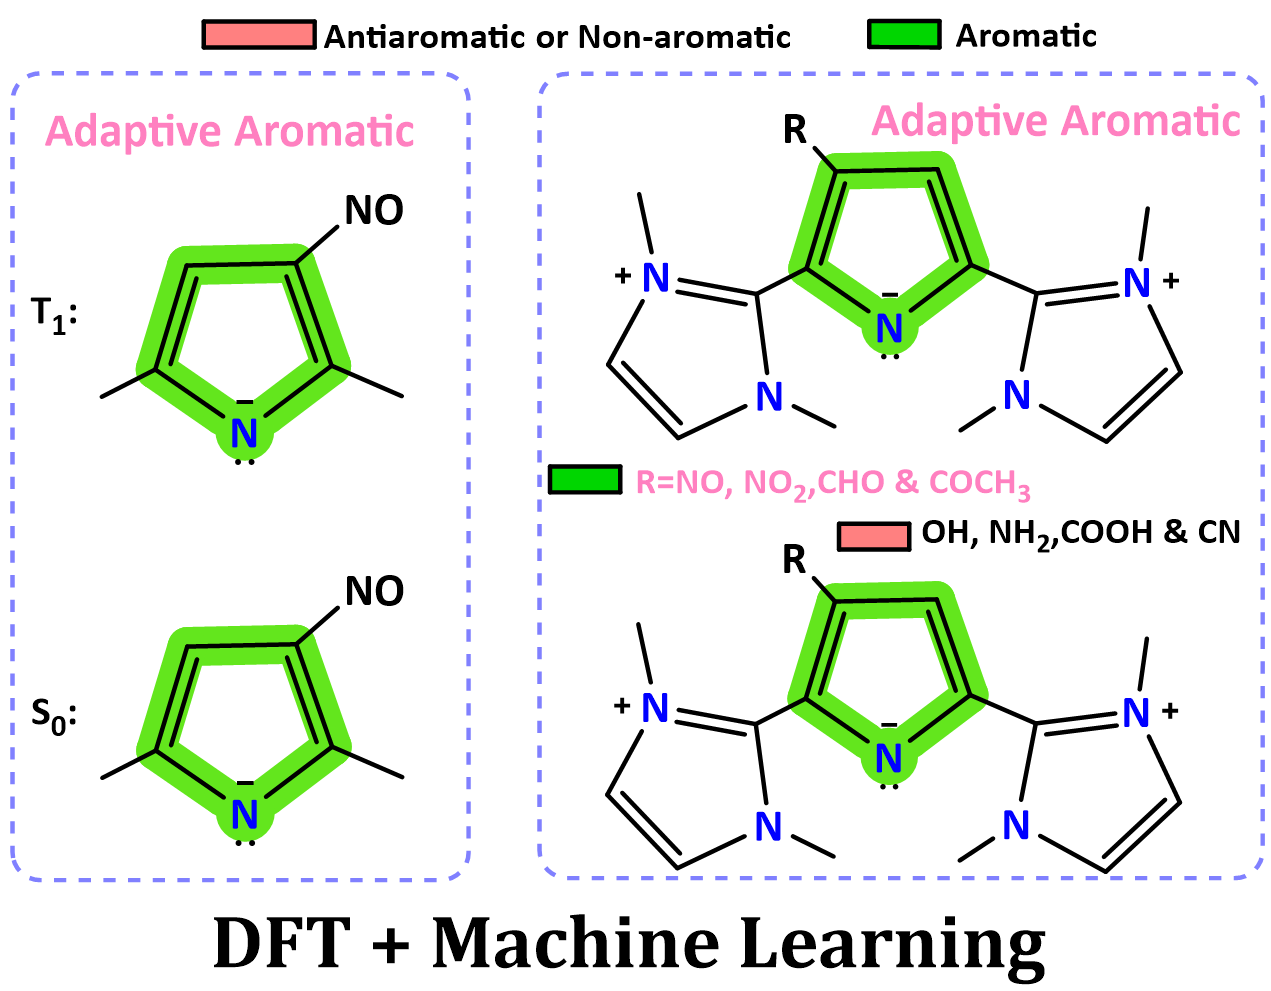
The stabilization of 4π-electron systems remains a fundamental challenge in chemistry, stemming from their intrinsic antiaromaticity and thermodynamic instability as dictated by Hückel’s rule. A recent experimental study has demonstrated that rational molecular design using N-heterocyclic carbenes (NHCs) can surmount these limitations by inducing aromaticity. However, the issue of T1 aromaticity in such systems remains unresolved. Elucidating the substituent-modulated behavior of T1 (anti)aromaticity is crucial for deciphering the physicochemical properties of these systems.

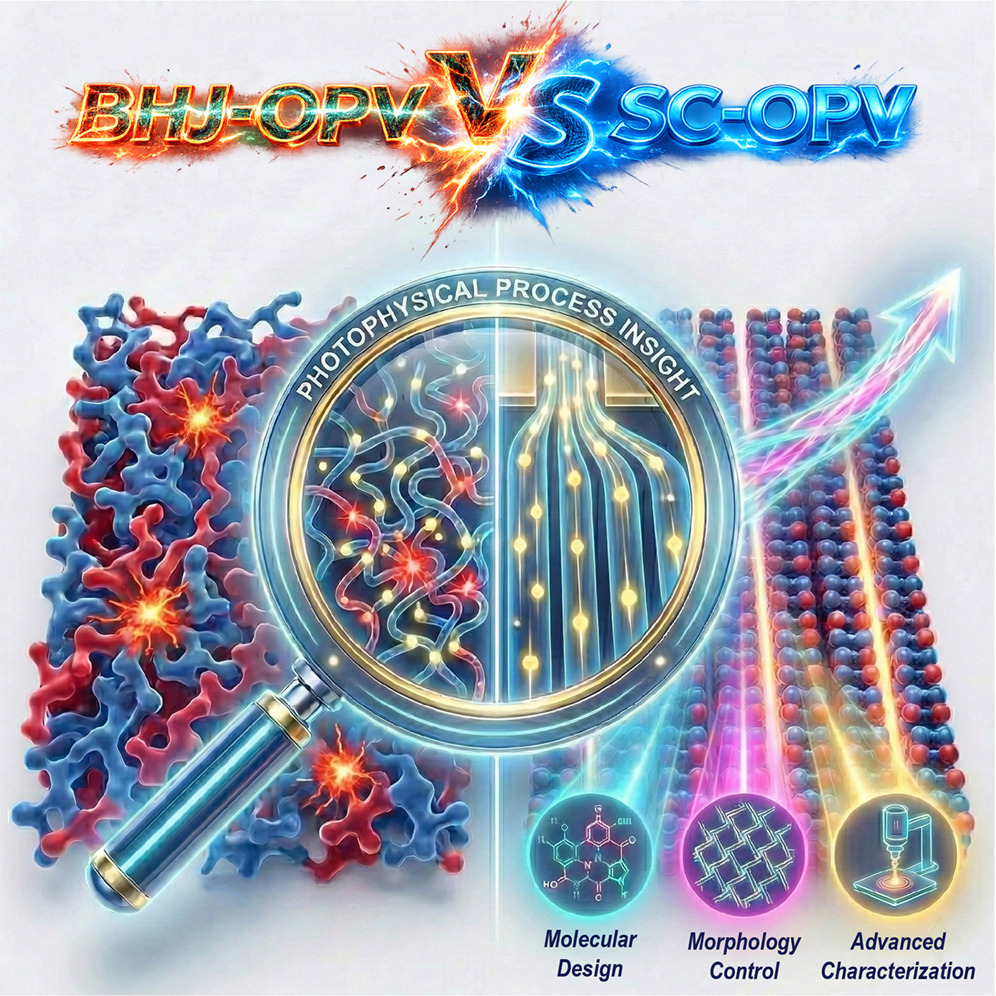
Single-component organic photovoltaics (SC-OPVs) offer an innovative approach to the traditional bulk-heterojunction organic photovoltaics (BHJ-OPVs) by chemically bonding the donor and acceptor materials within a single molecule. This approach has the potential not only to reduce the manufacturing costs but also to stabilize the photoactive layer against common morphological degradation processes typically observed in BHJ-OPVs.

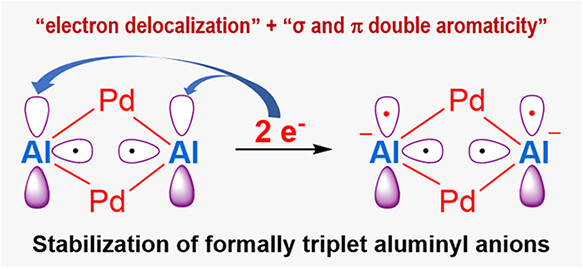
While triplet carbenes have been stabilized through spin delocalization, capturing triplet aluminyl anions─isoelectronic counterparts of carbenes─has remained an unsolved challenge in main-group chemistry. All previously reported aluminyl anions exhibit a singlet ground state, despite the use of diverse ligand frameworks. Herein, we report the first example of formally triplet aluminyl anions within bimetallic clusters, [((CH3)2C(CH2NPiPr2)2)2Al2Pd2]M2 (M = Li, Na, K).

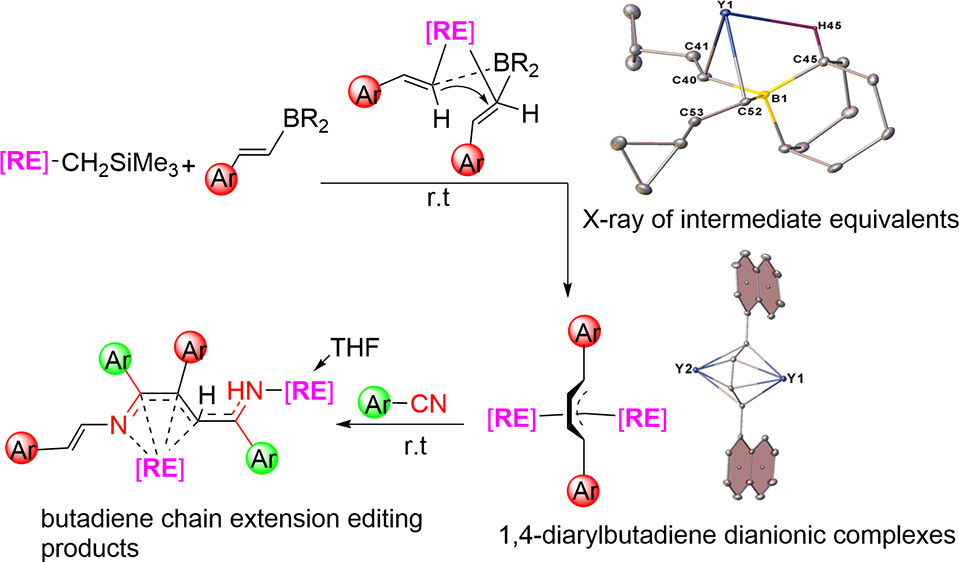
Aryl and vinyl boron reagents are widely employed in modern organic synthesis. However, the direct coupling of nucleophilic vinyl boron reagents to construct conjugated 1,4-diarylbutadiene dianionic (DABDA) motifs in the absence of additives remains a significant challenge. Here, we report for the first time the direct coupling of various β-aryl vinyl boron reagents, initiated by the rare-earth metal monoalkyl complexes for the synthesis of a number of rare-earth metal complexes featuring inverse sandwich DABDA motifs.
Copyright © 2026,
Theme Originally Created by Devsaran